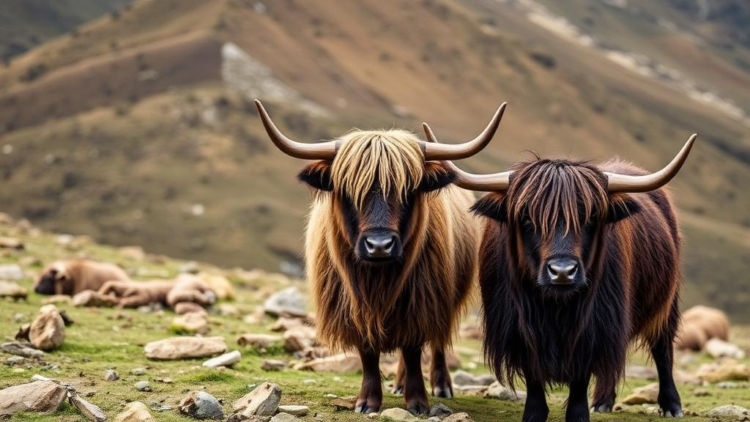
High on the Tibetan Plateau, where oxygen is scarce, winters are brutal, and ultraviolet radiation beats down unfiltered, lives one of the most remarkable livestock species on Earth. The yak is not merely a beast of burden for the herders of the region; it is a keystone of the entire high-altitude ecosystem and a living laboratory for studying how mammals adapt to extreme environments. Now, a team of Chinese researchers has taken one of the most detailed genomic looks yet at three little-known yak populations from the Xizang region, and their findings are reshaping how scientists understand the evolutionary history and breeding potential of this iconic animal.
The study, published in BMC Genomics, focused on three yak populations recently identified in the Xizang region as carrying unique germplasm characteristics: the Chawula yak, abbreviated CWL, the Jila yak, JL, and the Niangya yak, NY. Although these groups had attracted attention for distinctive traits, their genomic foundations and their relationships to other yak breeds remained poorly resolved. To close that gap, the researchers turned to whole-genome resequencing, a technique that reads the entire genetic blueprint of individuals rather than sampling only selected markers, allowing an extraordinarily fine-grained view of variation across the genome.
Whole-genome resequencing works by breaking each animal’s DNA into millions of short fragments, sequencing them, and then aligning the resulting reads against a reference genome. Where an individual’s sequence differs from the reference, researchers can call single nucleotide polymorphisms, or SNPs, as well as insertions and deletions of DNA letters known as InDels. Because this approach captures variation across the whole genome rather than at a handful of loci, it provides the statistical power needed to reconstruct population histories, measure inbreeding, and detect the faint fingerprints of natural and artificial selection that accumulate over thousands of generations.
The team did not stop at the three new populations. By integrating their data with genomic information from fifteen previously characterized yak populations, they built a comprehensive framework placing CWL, JL, and NY within the broader evolutionary landscape of Tibetan yak. This comparative design matters enormously in population genetics: diversity and differentiation statistics are only meaningful in context, and phylogenetic trees built from a handful of breeds can be misleading. With eighteen populations in the analysis, the researchers could ask whether the new groups represent genuinely distinct lineages or simply geographic extensions of known ones.
The answer surprised them. The Chawula, Jila, and Niangya yaks turned out to harbor richer genetic diversity than many of their counterparts, along with unique ancestral compositions and a moderate level of genetic differentiation from other yak populations. In practical terms, these animals carry a deep reservoir of genetic variation, which is exactly the raw material that breeding programs need when selecting for traits such as disease resistance, meat quality, or environmental tolerance. Populations with depleted diversity, by contrast, often struggle to respond to changing conditions, whether those changes are climatic, epidemiological, or market-driven.
Perhaps the most striking result came from the phylogenetic and structure analyses. The three new populations clustered together with the Yushu yak and wild yak into a distinct subclade, while the other domestic yak populations formed a separate cluster. Within that subclade, the relationships grew even more interesting: the Chawula and Niangya yaks were most closely related to each other, while the Jila yak grouped with the Yushu yak, a population that itself carries wild yak ancestry. This pattern suggests that the genetic history of Xizang yaks is more tangled, and more ancient, than a simple story of isolated herds diverging over time would predict.
Indeed, the researchers found that the genetic structure of yak populations in the Xizang region, including the three new groups, does not map neatly onto geography. Instead, the data revealed complex patterns of gene flow between populations, meaning that animals, and their genes, have moved across the plateau in ways that do not follow a straightforward distance-decay model. For conservation biologists, this is a crucial insight. It implies that managing yak genetic resources requires understanding historical exchange networks and ancestral contributions, not just drawing circles on a map around where each population currently lives.
Beyond reconstructing history, the study used selection signal analysis to hunt for genes shaped by selective pressures. The researchers combined two complementary statistics: F_ST, which measures the degree of genetic differentiation between populations at each point in the genome, and Pi, a measure of nucleotide diversity within populations. When both metrics flag the same genomic regions, those regions become strong candidates for having experienced selection. This dual-statistic approach helps filter out neutral demographic noise, such as the genome-wide effects of population bottlenecks, and home in on loci where something biologically interesting has happened.
The candidate genes that emerged span an intriguing range of functions. Several, including KIF27, MC1R, and SLC24A5, are potentially involved in coat color formation. MC1R is a familiar name in pigmentation genetics, famous across mammals from mice to humans for its role in switching between eumelanin, the dark pigment, and pheomelanin, the reddish one. SLC24A5 likewise has a celebrated pedigree, having been implicated in pigmentation differences in zebrafish and humans. Finding these genes under selection in yak populations with distinctive coat colors suggests a shared mammalian toolkit at work on the plateau. Other candidate genes, among them CP, ARPC2, and ATP1A2, point toward meat production performance, a trait of obvious economic importance for herding communities.
The pathway-level analysis added further depth. The researchers identified key biological pathways related to oxytocin signaling, melanogenesis, neural development, growth and development, digestion and absorption, and immune regulation. Each of these categories makes intuitive sense for an animal living at extreme altitude: efficient digestion and nutrient absorption matter when forage is sparse and seasonal, immune regulation matters in a harsh pathogen landscape, and growth pathways shape the body conformation that determines survival and productivity. The oxytocin pathway is particularly thought-provoking, given its roles in social bonding, reproduction, and stress responses, all of which could be under pressure in animals managed in traditional herding systems.
What makes this study more than an academic exercise is its practical payoff. By demonstrating at the genomic level that the Chawula, Jila, and Niangya yaks possess unique genetic backgrounds and considerable breeding potential, the research provides a scientific foundation for conservation and sustainable utilization of yak genetic resources. The candidate genes identified offer concrete targets for future selective breeding programs, potentially allowing herders to improve meat yield or fix desirable coat color traits without resorting to crossbreeding that could dilute the local adaptations these populations have accumulated. The results also lay the groundwork for functional validation studies, in which the roles of specific genes can be tested experimentally rather than inferred from population statistics alone.
There is also a broader scientific lesson here. The yak joins a growing roster of high-altitude species, from Tibetan humans to Tibetan mastiffs, whose genomes are teaching biologists how life copes with hypoxia, cold, and intense radiation. Every newly characterized population adds resolution to that picture, and the discovery that Xizang yaks form a distinct subclade with wild yak ancestry hints that untapped adaptive variation may still be hiding in the plateau’s remnant wild herds and semi-wild populations. Preserving that variation before it disappears, whether through habitat change, unmanaged hybridization, or the genetic erosion that accompanies small population sizes, is a race against time that genomics is helping to inform.
The work, led by researchers at the Sichuan Qinghai Tibet Plateau Herbivore Livestock Engineering Technology Center of Southwest Minzu University and the Xizang Academy of Agricultural and Animal Husbandry Sciences in Lhasa, was conducted under the Second Tibetan Plateau Scientific Expedition and Research Program, a major national effort to catalog and understand the plateau’s biological resources. As sequencing costs continue to fall and analytical methods grow more sophisticated, studies of this kind are likely to multiply, transforming our understanding of how domestic animals and their wild relatives have co-evolved with one of the planet’s most demanding environments. For the yak, the shaggy engine of Tibetan life, the genome is finally beginning to give up its secrets, and those secrets may prove as valuable to the herders of tomorrow as the animal itself has been to the herders of the past.
Subject of Research: Whole-genome resequencing of Tibetan yak populations to assess genetic diversity, population structure, and selection signatures
Article Title: Whole-genome resequencing reveals genetic diversity, population structure, and selection signatures in tibetan yak
Article References: Zhe, Y., Jiang, H., Zhang, Y., Sibinuer, Y., Shan, S., Wu, Z., Chai, Z., & Xin, J. (2026). Whole-genome resequencing reveals genetic diversity, population structure, and selection signatures in tibetan yak. BMC Genomics. https://doi.org/10.1186/s12864-026-13384-5
Image Credits: AI Generated
DOI: 10.1186/s12864-026-13384-5
Keywords: yak, whole-genome resequencing, genetic diversity, population structure, selection signatures, Tibetan Plateau, conservation genomics, MC1R, coat color, high-altitude adaptation, SNP, phylogenetics
Cite Scienmag News
APA MLA Chicago
Copy citation Download RIS
Tags: adaptation to extreme environmentscoat colorcomparative yak population studyconservation genomicsGenetic diversitygenetic diversity in Tibetan yaksgenomic insights into yak conservationhigh-altitude adaptationhigh-altitude livestock geneticslivestock breeding potentialMC1Rphylogeneticspopulation structureregional yak breed differentiationselection signaturesSNPTibetan PlateauTibetan Plateau animal adaptationunique yak germplasm characteristicswhole-genome resequencingwhole-genome resequencing in yaksyakyak evolutionary historyYak genomics